Disclaimer: Early release articles are not considered as final versions. Any changes will be reflected in the online version in the month the article is officially released.
Volume 32, Number 10—October 2026
Dispatch
Whole-Genome Sequencing of Measles Virus from 2025 Outbreak, Texas, USA
 , Bonnie Oh, Richard Bovio, Jie Lu, Sydney Stanley, Robert F. Potter, Yan Sun, Casey Schroeder, Olivia A. Smith, Elise Huebner, Stephen L. White, Saroj Rai, Varun Shetty, and Grace Kubin
Figure 2
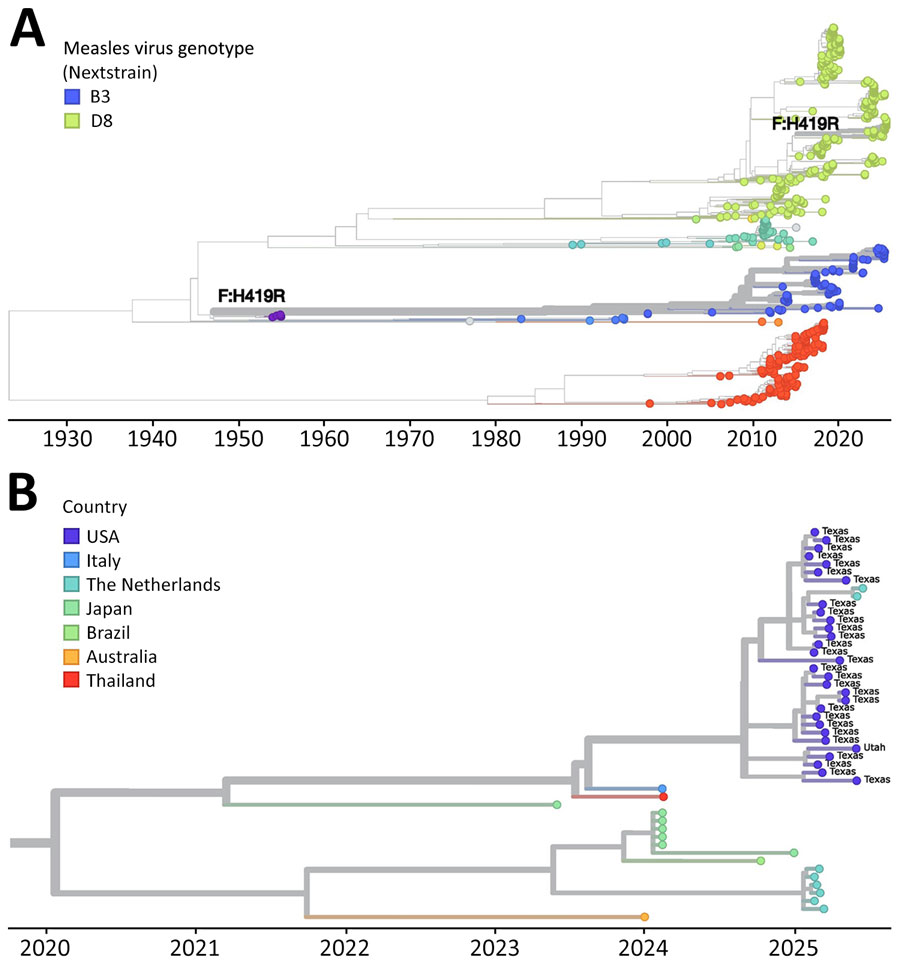
Figure 2. Genomic surveillance context of F:H419R among publicly available measles virus genomes from study of whole-genome sequencing of measles virus from 2025 outbreak, Texas, USA. A) Nextstrain time-scaled global phylogeny of measles viruses showing genotype B3 and D8 measles virus genomes with the F:H419R substitution highlighted (https://nextstrain.org/measles/genome@2025-09-24?gt=F.419R&treeZoom=selected). B) Global genotype D8 context of representative measles genomes associated with the outbreak in Texas and representative publicly available F:H419R-positive D8 genomes. F:H419R, Fusion gene H419R substitution.