Skip directly to site content Skip directly to page options Skip directly to A-Z link Skip directly to A-Z link Skip directly to A-Z link

Disclaimer: Early release articles are not considered as final versions. Any changes will be reflected in the online version in the month the article is officially released.

Volume 32, Number 10—October 2026

Dispatch

Whole-Genome Sequencing of Measles Virus from 2025 Outbreak, Texas, USA

Piyada JuntawongComments to Author , Bonnie Oh, Richard Bovio, Jie Lu, Sydney Stanley, Robert F. Potter, Yan Sun, Casey Schroeder, Olivia A. Smith, Elise Huebner, Stephen L. White, Saroj Rai, Varun Shetty, and Grace Kubin
Author affiliation: Texas Department of State Health Services, Austin, Texas, USA (P. Juntawong, B. Oh, R. Bovio, J. Lu, Y. Sun, C. Schroeder, O.A. Smith, E. Huebner, S.L. White, S. Rai, V. Shetty, G. Kubin); Children's Hospital of Philadelphia, Philadelphia, Pennsylvania, USA (S. Stanley, R.F. Potter); Perelman School of Medicine at the University of Pennsylvania, Philadelphia (S. Stanley, R.F. Potter)

Main Article

Figure 3

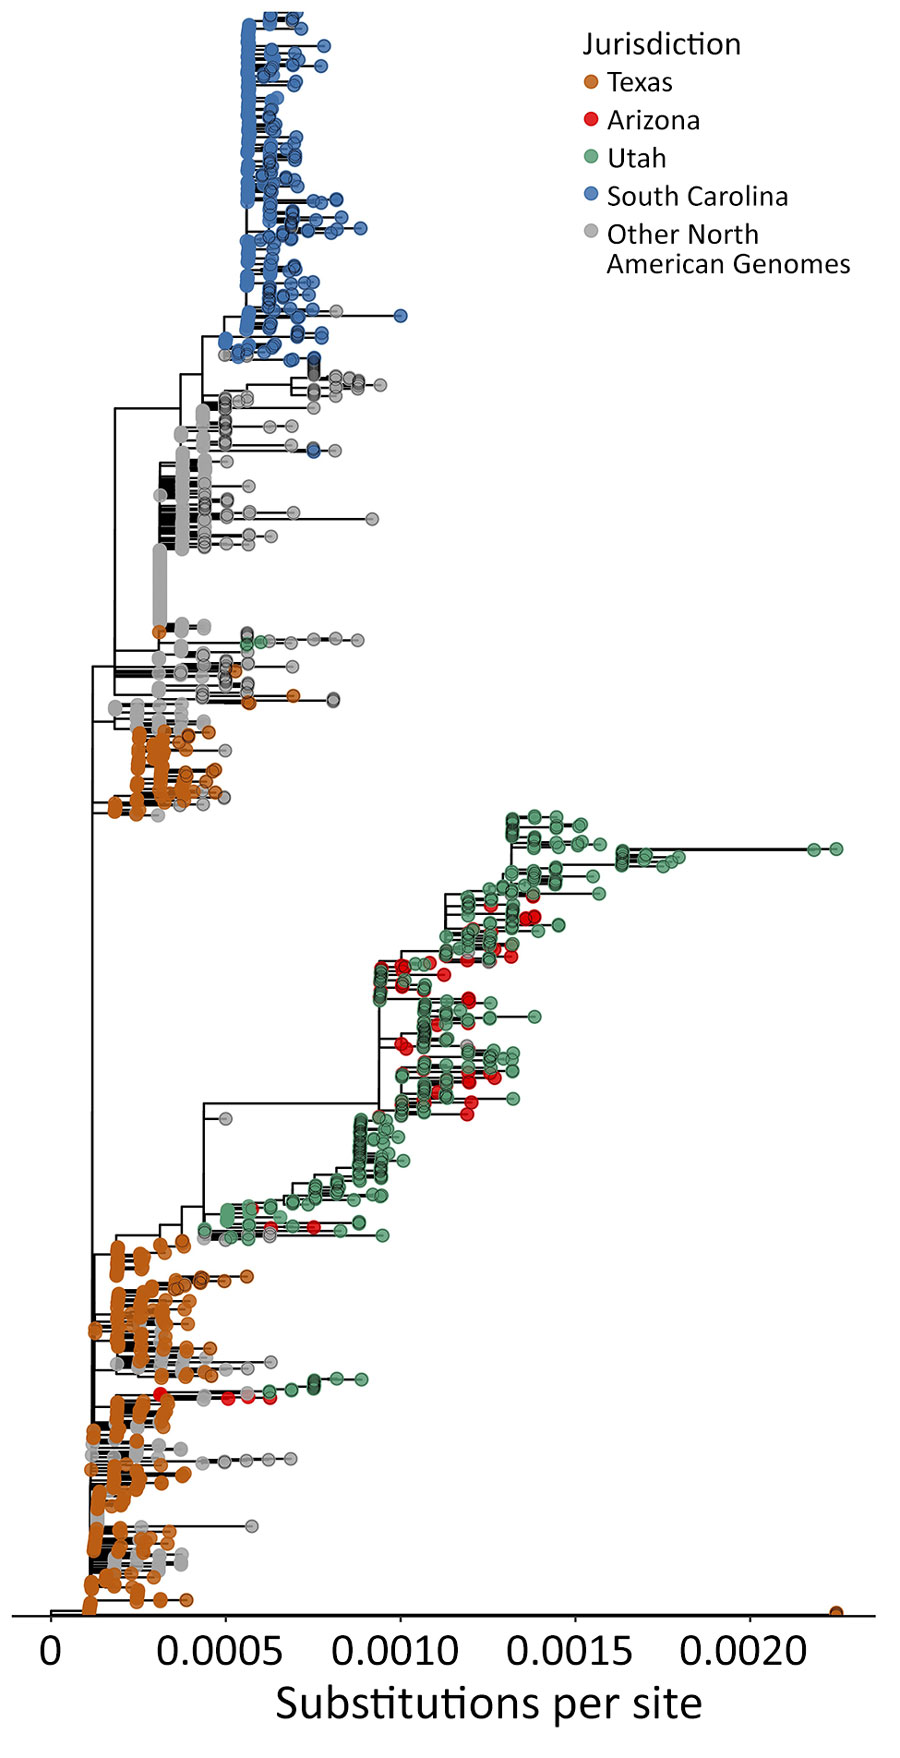
Maximum-likelihood phylogeny of the focal expanding North American genotype D8 measles virus lineage from study of whole-genome sequencing of measles virus from 2025 outbreak, Texas, USA. Phylogeny shows 1,760 genomes from Texas, Utah, Arizona, South Carolina, and other jurisdictions in North America included in the focal expanding North American genotype D8 lineage. We excluded 11 contextual genomes outside the focal lineage from this display. Tips are colored by jurisdiction; Canada genomes and US genomes outside Texas, Arizona, Utah, and South Carolina are grouped as other North American genomes. Tips identified by TreeTime (https://github.com/neherlab/treetime) as molecular-clock outliers are shown with black open circles. Branch lengths represent substitutions per site. The tree was rooted for visualization only; root placement should not be interpreted as evidence of ancestry, source, or transmission direction.

Figure 3. Maximum-likelihood phylogeny of the focal expanding North American genotype D8 measles virus lineage from study of whole-genome sequencing of measles virus from 2025 outbreak, Texas, USA. Phylogeny shows 1,760 genomes from Texas, Utah, Arizona, South Carolina, and other jurisdictions in North America included in the focal expanding North American genotype D8 lineage. We excluded 11 contextual genomes outside the focal lineage from this display. Tips are colored by jurisdiction; Canada genomes and US genomes outside Texas, Arizona, Utah, and South Carolina are grouped as other North American genomes. Tips identified by TreeTime (https://github.com/neherlab/treetime) as molecular-clock outliers are shown with black open circles. Branch lengths represent substitutions per site. The tree was rooted for visualization only; root placement should not be interpreted as evidence of ancestry, source, or transmission direction.

Main Article

Page created: September 10, 2026
Page updated: September 21, 2026
Page reviewed: September 21, 2026
The conclusions, findings, and opinions expressed by authors contributing to this journal do not necessarily reflect the official position of the U.S. Department of Health and Human Services, the Public Health Service, the Centers for Disease Control and Prevention, or the authors' affiliated institutions. Use of trade names is for identification only and does not imply endorsement by any of the groups named above.
file_external